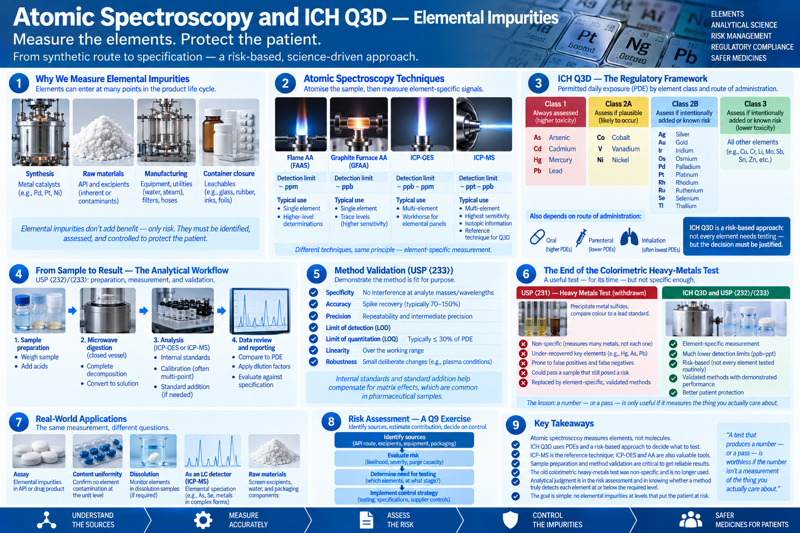
(Lecture 5.) NMR was MAI’s the last two weeks; this week the AF/SM thread picks back up where Week 2 left it, with the first of three technique weeks — atomic spectroscopy, then molecular spectroscopy, then separations and mass spectrometry. The API-synthesis section flagged metal catalysts as a source of elemental impurities. This week is where that risk gets measured, and where it either clears a limit or has to be controlled.
The one idea
A test that produces a number — or a pass — is worthless if the number isn’t a measurement of the thing you actually care about.
Atomic spectroscopy — the technique family
Where UV-Vis measures molecules, atomic spectroscopy measures elements: atomise the sample, then measure absorption or emission at element-specific wavelengths, or count ions by mass.
| Technique | Detection | Typical use |
|---|---|---|
| Flame AA | ppm | Single-element, higher-level (e.g. residual catalyst at limit) |
| Graphite furnace AA (GFAA) | ppb | Single-element trace |
| ICP-OES | ppb–ppm, multi-element | Workhorse for panels of elements |
| ICP-MS | ppt–ppb, multi-element, isotopic | Trace elemental impurities; the Q3D reference technique |
The regulatory frame — ICH Q3D / USP ⟨232⟩–⟨233⟩
Q3D sets permitted daily exposures (PDEs) for elemental impurities by route of administration, grouped into classes: Class 1 (As, Cd, Hg, Pb — always assessed), Class 2A (Co, V, Ni — likely, assess if plausible), Class 2B and Class 3 (assessed only if intentionally added, such as a catalyst named in the synthesis, or otherwise a known risk). The analyst’s job splits in two:
- The risk assessment — where could each element come from (drug-substance synthesis and catalysts — the API-synthesis section’s Pd, Pt, Ni couplings, for instance), excipients, water, manufacturing equipment, container closure — and does the total plausibly approach the PDE? This is a Q9 risk-management exercise, and it decides whether routine testing is even needed.
- The method — closed-vessel microwave digestion, then ICP-OES or ICP-MS, validated per USP ⟨233⟩ (specificity, accuracy by spiked recovery, precision, a demonstrated limit) with internal standards and often standard addition for matrix effects.
Worked case — from “heavy metals” to element-specific testing
Until the 2010s, most pharmacopeias controlled elemental impurities with a single colorimetric heavy-metals test (the old USP ⟨231⟩): precipitate metal sulfides, compare the resulting brown colour to a lead standard, report pass/fail. It was cheap and universal — and wrong in both directions. It under-recovered the elements of most concern (mercury, and much of the arsenic and lead, were lost in sample preparation) and it flagged samples on colour that had nothing to do with toxic metals. ICH Q3D and USP ⟨232⟩/⟨233⟩ replaced it with a risk assessment plus element-specific, validated instrumental methods. The lesson is specificity: a test that produces a number, or a pass, is worthless if the number isn’t a measurement of the thing you care about.
Risk-assessment assignment (Risk Homework, checkpoint 1 of 3)
Using the FMEA / risk-ranking approach from Week 2: take a drug product of your choice and build the Q3D elemental-impurity risk assessment — sources (starting with the synthetic route), contributions, and a defended conclusion on which elements, if any, need routine testing and at what stage. The same assignment structure returns at molecular spectroscopy and mass spectrometry — by the third pass, scoring detectability should be a habit.
Where the analyst sits
Q3D makes elemental impurities feel like a checklist — classes, PDEs, a table of elements. The judgment is upstream of the table: does the method referenced in a risk assessment actually detect each named element at 30% of its PDE, or is the assessment leaning on a method that was never challenged to see that low? Checking that is the job, not trusting that someone already did.
For discussion
- The old colorimetric heavy-metals test had a %RSD better than many ICP methods. Why is that not reassuring?
- A Q3D risk assessment concludes “no routine testing needed” for palladium, based on the synthetic route in the API-synthesis section. What evidence would you want to see before accepting that conclusion?
- If your ICP-MS method has a validated limit of quantitation right at 30% of an element’s PDE, is that method fit for purpose? What would make you more or less comfortable with it?
Source note. Elemental impurities: ICH Q3D(R2), USP ⟨232⟩/⟨233⟩, and the history of the withdrawn ⟨231⟩.